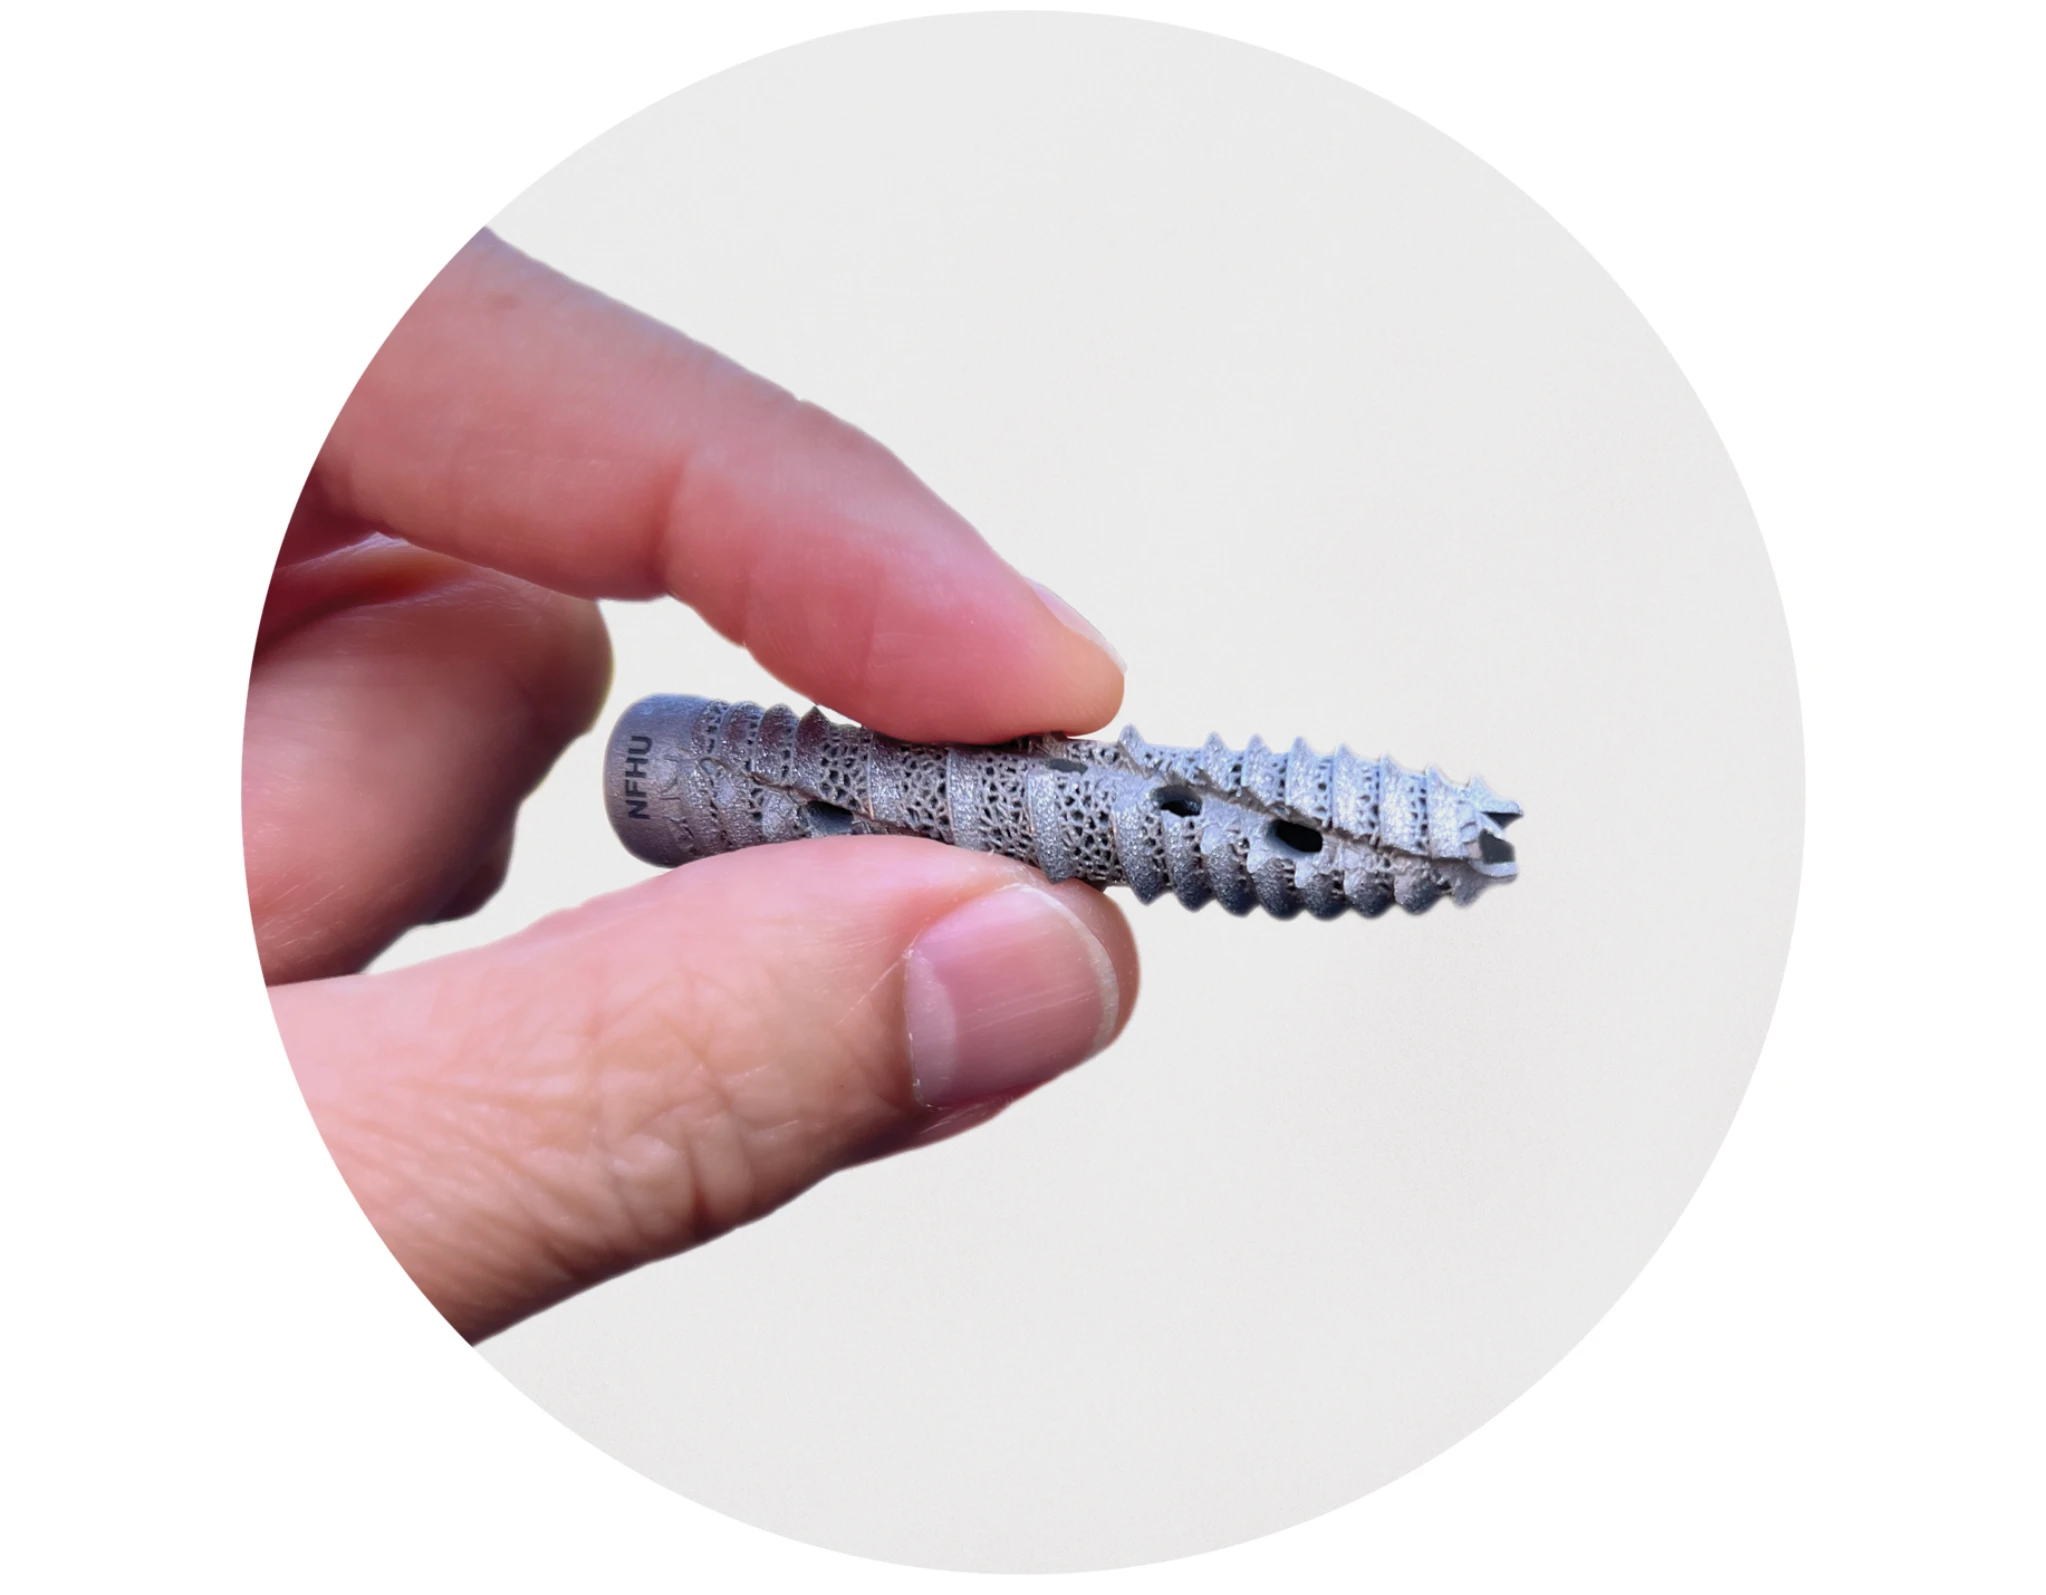
Arrowhead DE Arrow-LOK Implant
The Arrow-LOK Implant did not undergo required gamma radiation dose audit, which could lead to patient infection.
Moderate risk
Using it may cause temporary or medically reversible health problems. Serious harm is unlikely.
FDA Class II recall · How risk is rated
What happened
Arrowhead DE LLC of Peachtree City, Georgia, recalled the Arrow-LOK Implant, used for fixation of osteotomies, arthrodeses and reconstruction in the lesser toes following corrective procedures, because the implant did not undergo required gamma radiation dose audit. A potential reduction in sterility assurance could lead to patient infection.
The company began the recall on June 23, 2026, and it was added to the FDA's Enforcement Report on August 14, 2026. Distribution was nationwide in the states of VT, FL, TN, NY, GA, TX, and HI.
Summary by Recall Canary from the FDA notice.
What to do
We don't have product-specific instructions for this recall. FDA's general advice for medical device recalls:
- A device recall doesn't always mean you must stop using it: it may need to be checked, adjusted or fixed.
- Talk to your health care provider before you stop using it.
Is yours affected?
Affected if its UDI-DI and lot code are listed under that product.
Arrow-LOK Implant, REF
- UDI-DI
- 00855967006035
- 24662119
- 00855967006042
- 24662122
- 00855967006059
- 24662125
- 00855967006066
- 24662128
- 00855967006073
- 00855967006080
- 24662133
- 00855967006097
- 24662135
- 00855967006110
- 00855967006127
- 00855967006134
- 24662150
- 00855967006004
- 24662113
- 00855967006011
- Lot
- 21-253519
- 242162
Details
- Company began the recall
- Sold in
US Nationwide distribution in the states of Vermont, Florida, Tennessee, New York, Georgia, Texas, Hawaii.
- Event ID
- 99383
- Recall No.
- Z-2958-2026
- Company
- Arrowhead DE LLCPeachtree City, Georgia
- Hazard
- Sterility failure
- Recall type
- Voluntary: Firm initiated
- Customers told by
- Two or more of the following: Email, Fax, Letter, Press Release, Telephone, Visit
Company began the recall
Added to FDA's Enforcement Report · Classified by FDA
Listed on Recall Canary
In FDA's weekly enforcement report
Summary updated