FDA recall, , Moderate risk
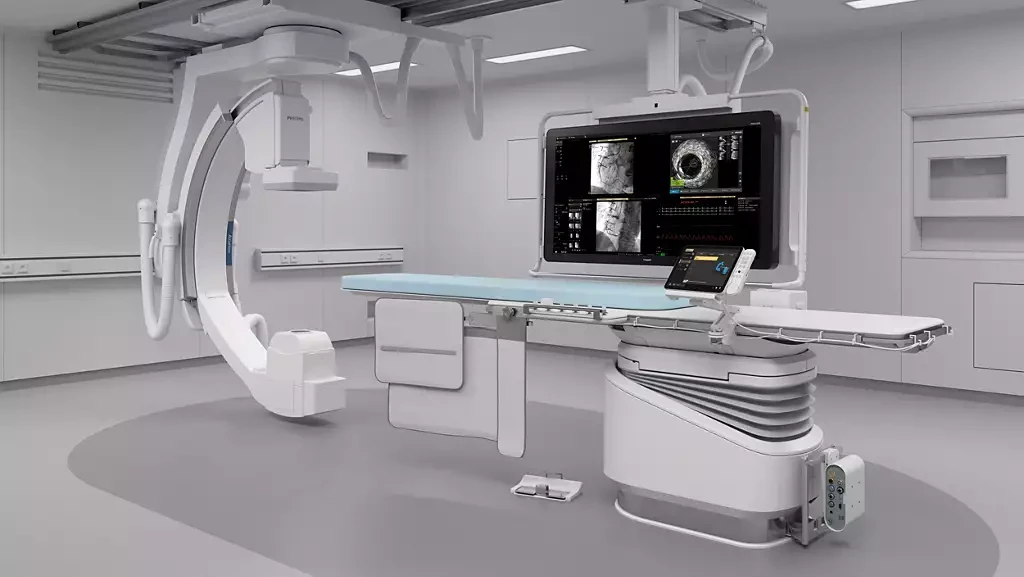
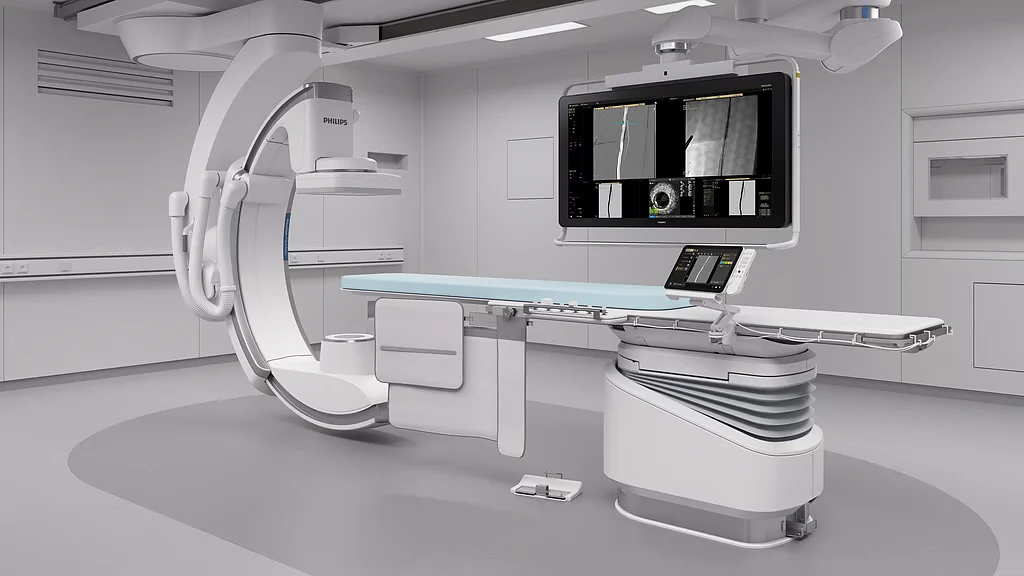
Philips Azurion 3, 5 and 7 Systems
Philips Azurion 3, 5 and 7 Systems: the Power Distribution Unit does not provide direct current power to essential components during a start or restart.
What to do
Check yoursIf a code on your device is listed below:
- Check the lot against the recall notice and ask your provider or supplier before you stop using it.
Typical advice for a recalled medical device; the FDA notice gives no product-specific instructions.

Summary by Recall Canary from the FDA notice
What happened
Philips Medical Systems Nederland B.V. is recalling Philips Azurion systems. When a power supply failure occurs during a start or restart, the Power Distribution Unit does not provide the required direct current power to essential system components. As a result, the system does not complete the start or restart, imaging functionality is unavailable, and the system remains unavailable for clinical use.
The recall covers 2,632 units of Philips Azurion 3, 5 and 7 systems, including the Azurion 3 M12, Azurion 5 M20 and Azurion 7 B20 models. The company started the recall on its own on October 15, 2026, and told customers by letter. The products were distributed nationwide in the United States and in many other countries. The FDA classified the recall on November 18, 2026.
Is yours affected?
Match the list below, then check the codes the FDA notice also names.
Philips Azurion systems
Product details from the notice
Philips Azurion systems. Containing the below descriptions and corresponding Model Numbers. 1. Azurion 3 M12, Model Numbers: 722063, 722221. 2. Azurion 3 M15, Model Numbers: 722064, 722222. 3. Azurion 5 M12
- Model
- 722227
Recall details
- Posted by FDA
- Recall began
- Status
- Open
- Weekly archive
- Listed the week of Nov 16, 2026
- Sold in
Distributed nationwide and in Afghanistan, Albania, Algeria, Argentina, Australia, Austria, Bangladesh, Belgium, Bolivia, Brazil, Brunei Darussal, Bulgaria, Canada, Chile, China, Colombia, Costa Rica, Croatia, Cyprus, Czech Republic, Denmark, Ecuador, Egypt, Estonia, Finland, France, French Polynesia, Germany, Ghana, Greece, Hong Kong, Hungary, India, Indonesia, Iran, Iraq, Ireland, Israel, Italy, Japan, Jordan, Kazakhstan, Kenya, South Korea, Kosovo, Kuwait, Lao, Latvia, Lebanon, Lithuania, Malaysia, Mexico, Mongolia, Morocco, Myanmar, Nepal, Netherlands, New Zealand, Nicaragua, Nigeria, Norway, Oman, Pakistan, Palestine, Panama, Peru, Philippines, Poland, Portugal, Puerto Rico, Reunion, Romania, Russia, Saudi Arabia, Serbia, Singapore, Slovakia, Slovenia, South Africa, Spain, Sri Lanka, Sweden, Switzerland, Syria, Taiwan, Thailand, Tunisia, Turrkiye, Turkmenistan, Ukraine, United Kingdom, United Arab Emirates, Uzbekistan, Vietnam and Yemen.
- Recalled by
- Philips Medical Systems Nederland B.V.Best, Netherlands
- Hazard
- Device defect
- Risk
Moderate risk
Using it may cause short-term health problems, or ones that treatment can reverse. Serious harm is unlikely.
FDA Class II recall · How risk is rated
- FDA event number
- 100086
- Recall number
- Z-0482-2027
- Quantity recalled
- 2,632 units
- Recall type
- Started by the company
- How the company first told customers
- Letter
All recalls from Philips Medical Systems Nederland B.V.
Timeline
FDA's Enforcement Report is its public list of recalls; FDA posts a recall by adding it there.
PHILIPS MEDICAL SYSTEMS NEDERLAND B.V. began the recall
FDA posted the recall to its Enforcement Report and rated it Class II (moderate risk)
Added to Recall Canary